

What Makes GenomeBeans Different from Traditional NGS Tools?
Key Takeaways
- Traditional NGS tools demand coding and heavy infrastructure.
- GenomeBeans offers a no-code, cloud-first platform.
- Standardized pipelines ensure reproducibility.
- Built-in variant visualization accelerates insights.
- Collaboration tools simplify cross-lab teamwork.
- Supports RNA-seq, metagenomics, and variant calling in one platform.
- A practical alternative for labs without a dedicated bioinformatics team.

Next-Generation Sequencing (NGS) is central to modern biology, but analyzing raw data often remains challenging. Tools like SAMtools or GATK are powerful, yet they demand coding expertise, custom scripting, and high-end infrastructure—barriers for many researchers.
Choosing the right NGS data analysis tools directly affects your turnaround time, accuracy, and cost.
No-Code NGS Analysis With GenomeBeans
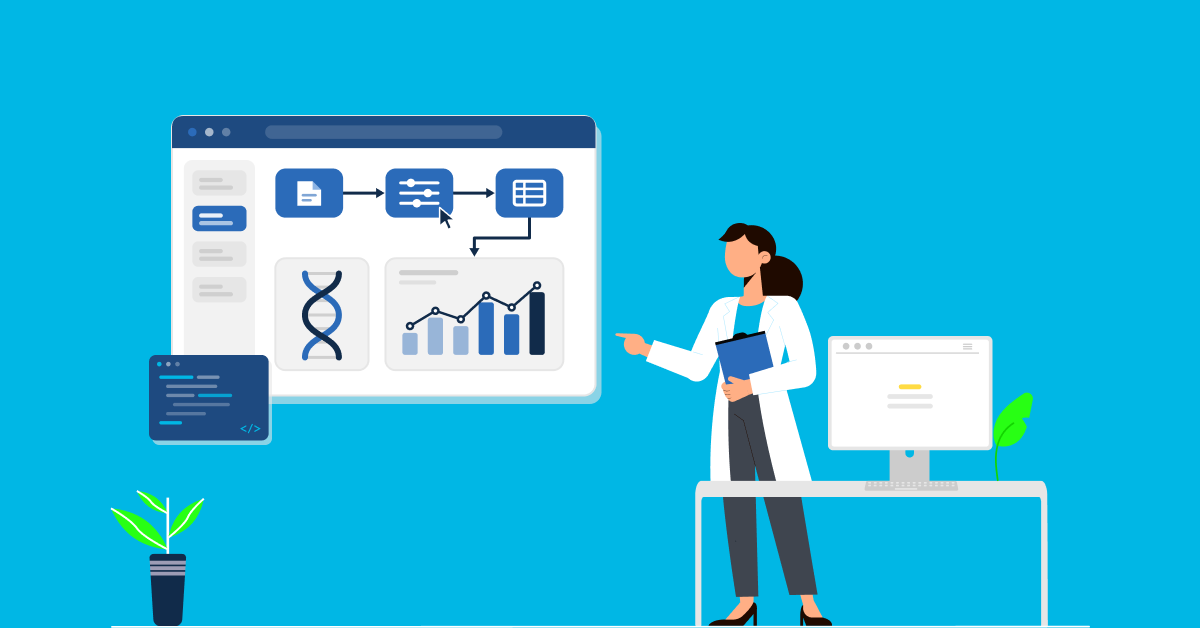
GenomeBeans eliminates these hurdles with a visual, no-code interface. Researchers can upload data, select workflows, and interpret results without programming. Being cloud-based, it also scales effortlessly without the need for local servers or IT support.
Wondering whether no-code NGS analysis can really replace scripting? It works best for routine RNA-seq, metagenomics, and variant workflows.
Standardized NGS Pipelines for Reproducibility
Traditional tools offer flexibility but risk inconsistencies across runs. GenomeBeans provides pre-validated, standardized pipelines for RNA-seq, metagenomics, and variant calling, ensuring reproducibility while still allowing controlled customization.
Pipelines are ready to use for RNA-seq, metagenomics, and variant calling, so teams get consistent results across runs.
Single Automated NGS Workflow
Instead of chaining multiple tools, GenomeBeans integrates quality control, alignment, quantification, and visualization into a single automated workflow. This reduces manual steps, errors, and time spent troubleshooting.
What this replaces:
- Separate QC, alignment, and quantification scripts.
- Manual file handling between tools.
- Version and dependency conflicts across environments.
Variant Visualization and TMB Analysis
GenomeBeans goes beyond standard workflows by offering variant-focused visualizations for example, identifying the top 10 mutated genes or summarizing tumor mutation burden (TMB). These insights make it easier for researchers to connect raw sequencing data with biological meaning and clinical relevance.
Collaboration for Research Teams
With project sharing, version history, and annotations, GenomeBeans supports cross-lab and interdisciplinary teamwork, streamlining research communication and ensuring transparency.
Quick Comparison: GenomeBeans vs Traditional NGS Tools
| Feature | Traditional Tools (SAMtools, GATK) | GenomeBeans |
|---|---|---|
| Coding Required | Yes | No (visual interface) |
| Infrastructure | Local servers / HPC | Cloud-based |
| Reproducibility | Depends on scripts | Standardized pipelines |
| Visualization | Separate tools needed | Built-in visualizations |
| Collaboration | Manual sharing | Project sharing and version history |
| Result Interpretation | Requires manual interpretation | Interpretive reports |
| Publication-Ready Figures | Often requires separate tools and manual formatting | Publication-ready figures |
Conclusion
GenomeBeans reimagines NGS workflows by making them accessible, reproducible, and collaborative while traditional tools remain powerful for highly customized use cases.
Frequently Asked Questions
1. Is GenomeBeans a good alternative to GATK and SAMtools?
For researchers who want results without coding or local servers, yes. GATK and SAMtools remain strong for highly customized pipelines.
2. Do I need programming skills for NGS data analysis on GenomeBeans?
No. You upload data, choose a workflow, and view results through a visual interface.
3. Which NGS analyses does GenomeBeans support?
RNA-seq, metagenomics, and variant calling, with variant visualizations such as top mutated genes and TMB.
Popular Post
-
16 November 2026
How to Choose Between IMGT, Kabat, and Chothia for Antibody Analysis: What Researchers Should Know
-
11 November 2026
How Scientists Track the Way Your Immune System Actually Responds
-
02 November 2026
Which Bioinformatics Platform Fits Your RNA-seq Research? A Feature-by-Feature Comparison
-
27 October 2026
The Hidden Microbial World: How Metagenomics Reveals Microbes Beyond What We Can See
-
19 October 2026
Can No-Code Bioinformatics Replace Coding? Here’s Where It Helps